What software does chemists use is a question that opens the door to a fascinating digital revolution quietly reshaping the landscape of modern science. Long before a single drop of liquid touches laboratory glassware, brilliant minds are already mapping out complex molecular destinies across glowing computer screens. This profound shift bridges meticulous academic research with industrial breakthroughs, transforming raw data into life-changing discoveries through the power of advanced computation.
Step inside today’s high-tech laboratories and you will find an intricate ecosystem where advanced digital platforms govern everything from quantum mechanical modeling to strict regulatory safety tracking. As researchers decode the language of atoms and reactions with unprecedented precision, integrated software suites act as the invisible backbone of innovation. They not only streamline the chaotic flow of experimental data but also safeguard intellectual property, empowering global scientific communities to push the boundaries of what is humanly possible.
Chemists rely on specialized computational platforms to model molecular behavior and predict chemical reactions accurately before touching any laboratory glassware.
Long before the hiss of Bunsen burners echoes through modern laboratories or the scent of organic solvents fills the air, contemporary chemistry begins in the silent, illuminated glow of computer monitors. Scientists manipulate digital atoms and construct complex molecular architectures with unprecedented precision, shifting the paradigm of discovery from empirical trial and error to targeted digital foresight. This proactive approach saves countless hours and valuable resources, transforming theoretical concepts into tangible breakthroughs safely within virtual space.
The journey of digital molecular modeling began in the mid-20th century as a modest endeavor relying on physical wire-and-plastic skeletal kits before transitioning to rudimentary mainframe computers that calculated basic geometric distances. Decades of exponential growth in processing power propelled these primitive wireframe displays into sophisticated graphical interfaces capable of rendering proteins in vivid, real-time three dimensions. Industrial pharmaceutical laboratories and academic research centers alike adopted these platforms to visualize macromolecular targets, moving from rigid static representations to dynamic simulations that mirror the chaotic, fluid reality of biological systems.
While molecular modelers rely on specialized computational software to map complex atomic bonds with vivid precision, modern data operations demand equally sharp digital tools. Professionals quickly extract emails from call data ending with hightouch.com or hightouch.io to streamline vital enterprise communications. Ultimately, mastering these diverse digital platforms empowers researchers to accelerate groundbreaking laboratory discoveries efficiently.
Quantum mechanical calculations in modern software environments, What software does chemists use
Executing quantum mechanical calculations within modern computational suites requires solving the fundamental equations governing subatomic particles, primarily the time-independent Schrödinger equation. Because analytical solutions are impossible for systems with more than one electron, software environments rely heavily on approximation methods such as Hartree-Fock theory and Density Functional Theory to navigate this mathematical complexity. Researchers input the spatial coordinates of atomic nuclei and the total charge of the system, establishing a baseline geometry that the software subjects to iterative energy minimization algorithms.
The program constructs a Hamiltonian operator representing the total energy of the electrons and nuclei, utilizing mathematical sets known as basis functions to describe atomic orbitals. As the software iterates through self-consistent field procedures, it calculates electron-electron repulsion and nuclear attraction until the energy change between successive cycles falls below a strict convergence threshold. This rigorous computational choreography yields a wealth of electronic properties, including electrostatic potential maps, dipole moments, and vibrational frequencies, which accurately forecast how a molecule will behave in an actual chemical environment.
Real-world applications, such as the rapid virtual screening of drug candidates during the global health crisis, highlight how these calculations allow researchers to pinpoint viable molecular binders in days rather than years. Advanced visualization modules then render these dense matrices of numerical output into intuitive, color-graded charge distributions that guide synthetic chemists precisely where to build or cleave chemical bonds.
Quantum mechanics transforms the invisible dance of electrons into predictable pathways for molecular innovation.
Modern electronic structure calculation programs integrate a diverse array of advanced mathematical algorithms to solve complex quantum mechanical equations efficiently. These underlying computational engines determine the reliability, speed, and accuracy of molecular simulations across various scientific disciplines.
- Self-Consistent Field algorithms iteratively solve the non-linear equations governing electron movement until the calculated electron density stops changing between cycles.
- Basis set expansions employ linear combinations of mathematical functions to approximate unknown molecular orbitals with high fidelity.
- Density Functional Theory approximations map the total energy of a system as a functional of the electron density, balancing computational speed with high accuracy for large molecular systems.
- Geometry optimization routines utilize analytical gradients of potential energy surfaces to navigate atoms toward their most stable, lowest-energy spatial arrangements.
- Numerical integration techniques divide continuous molecular space into discrete grid points to evaluate exchange-correlation energies accurately across complex electronic topographies.
Laboratory information management systems transform how physical samples and experimental results are tracked from initial synthesis through final publication.
Every breakthrough molecule starts as a fragile idea whispered inside a fume hood, eventually evolving into a tangible physical sample that must endure relentless scrutiny before reaching the world. Modern discovery thrives when chaotic benchtop chemistry meets the unyielding precision of digital architecture. In the high-stakes arena of modern science, losing track of a single vial can mean months of wasted synthesis and millions of dollars in delayed therapeutics.
The journey of a chemical compound from a nascent flask reaction to a peer-reviewed journal article demands an unbroken chain of custody. Advanced informatics platforms step into this chaotic landscape to act as the digital backbone of the modern laboratory, capturing every weigh-in, purity test, and spectral analysis with unflinching accuracy. This digital evolution ensures that human error is intercepted before it cascades into flawed research outcomes, safeguarding both scientific integrity and commercial viability.
Core architectural components required for maintaining rigorous audit trails in regulated pharmaceutical environments
Regulatory bodies such as the Food and Drug Administration demand absolute transparency, requiring every digital footprint to tell an unalterable story of creation, modification, and analysis. Achieving this level of compliance is not merely about storing files; it requires a sophisticated triad of architectural safeguards designed to withstand internal tampering and external cyber threats alike. These foundational pillars ensure that data integrity remains uncompromised throughout the entire lifecycle of a pharmaceutical asset.
The engineering of these compliant frameworks relies on immutable ledger technologies and time-stamping mechanisms that permanently bind user identity to every keystroke and instrument read. Below are the essential system layers that guarantee full regulatory compliance and operational transparency:
- Cryptographic hashing protocols that immediately flag any unauthorized backend database modifications or manual data tampering attempts.
- Granular event-logging engines that capture pre-state and post-state values for every single parameter altered within a given experimental run.
- Isolated staging environments where audit logs are replicated in real-time, preventing administrative override of historical records.
Data integrity in regulated spaces is achieved only when the effort to falsify a record vastly exceeds the effort to perform the science legitimately.
Instrument integration methods and centralized data repositories
Bridging the physical divide between analytical hardware and digital databases remains one of the most critical engineering feats in modern laboratory informatics. When a high-performance liquid chromatograph finishes a complex assay, the raw electrical signals must be translated into structured datasets without human transcription errors. Modern laboratories deploy a sophisticated blend of middleware agents, application programming interfaces, and standardized communication protocols to ingest physical metrics directly into centralized cloud repositories.
This seamless conduit eliminates the clipboard era of science, replacing manual data entry with lightning-fast machine-to-machine handshakes that preserve signal fidelity from detector to dashboard.
Consider the daily operations at a major pharmaceutical development center where dozens of mass spectrometers run around the clock. Without automated integration, technicians would spend hours manually exporting proprietary file formats, converting them, and uploading them into shared network drives—a process fraught with risks of misplaced decimal points and mislabeled sample IDs. Instead, modern middleware acts as a universal translator, intercepting raw output files straight from the instrument control PC, stripping away proprietary encryption layers, and parsing the critical values into standardized schemas like JCAMP-DX or customized JSON payloads.
These structured packets are then securely transmitted over encrypted channels to enterprise relational databases, where they trigger automated verification scripts. If a sample falls outside acceptable baseline tolerances, the system instantly flags the deviation, alerting quality assurance personnel via secure webhooks while simultaneously locking the associated instrument queue. This automated synchronization not only accelerates throughput by upwards of seventy percent but also creates a pristine, unbroken evidentiary chain that satisfies the strictest criteria enforced by global regulatory authorities.
Workflow parameters, security protocols, user permissions, and backup frequencies
Managing laboratory data effectively requires a delicate balance between open collaboration and ironclad security, ensuring that operational speed never compromises regulatory compliance. The following matrix Artikels the operational parameters required to maintain a secure, traceable, and resilient laboratory informatics ecosystem:
| Sample Tracking Workflow | Security Protocol | User Permission Level | Backup Frequency |
|---|---|---|---|
| Initial Reagent Intake and Barcoding | End-to-end AES-256 encryption at rest and in transit | Junior Technician (Read/Write assigned batches) | Continuous real-time replication |
| Mid-phase Synthesis and Reaction Monitoring | Multi-factor authentication with hardware tokens | Senior Chemist (Full execution and parameter tuning) | Hourly differential snapshots |
| Analytical Testing and Spectral Validation | Role-based access control with biometric verification | Quality Assurance Officer (Review and Sign-off only) | Daily full system backup |
| Final Archival and Regulatory Submission | WORM (Write Once, Read Many) optical storage | System Administrator (Audit management only) | Weekly off-site immutable vaulting |
Electronic laboratory notebooks replace traditional paper records by enabling secure digital documentation and seamless team collaboration across geographically diverse research facilities
Modern scientific discovery thrives on agility, pushing laboratories to abandon bound paper pages in favor of cloud-enabled ecosystems where brilliant minds from Tokyo to Boston instantly share breakthrough synthesis pathways. This digital transformation breathes fresh momentum into day-to-day experiments, turning static notebooks into dynamic hubs of innovation while safeguarding proprietary breakthroughs against modern digital vulnerabilities.
As research institutions cross this digital frontier, safeguarding highly sensitive intellectual property remains a paramount priority. Cryptographic timestamping mechanisms establish an immutable audit trail, ensuring that every molecular structure drawn and every reaction yield recorded receives a mathematically locked seal of authenticity. These cryptographic protocols prevent retroactive alterations, satisfying rigorous patent office standards worldwide and shielding billion-dollar chemical portfolios from industrial espionage or premature disclosures.
Intellectual property protection and cryptographic timestamping in digital archiving
Safeguarding digital assets demands robust, multi-layered defense frameworks that seamlessly integrate into the daily workflow of bench chemists without causing frustrating operational bottlenecks. Advanced cryptographic hashing functions, such as SHA-256, generate unique digital fingerprints for every experimental entry saved within the system architecture. When a researcher finalizes a synthesis report, the platform automatically anchors this hash to a secure blockchain or trusted timestamp authority, creating an unbreakable chronological chain of custody that withstands the most rigorous legal scrutiny during patent litigation proceedings.
Regulatory compliance heavily relies on these automated verification layers to maintain data integrity across distributed research teams operating in different legal jurisdictions. Encryption standards safeguard valuable trade secrets both in transit across cloud networks and at rest within decentralized server clusters, ensuring absolute confidentiality. Organizations implement strict Role-Based Access Control policies combined with multi-factor authentication to monitor precisely who views, edits, or authorizes specific chemical formulations.
This rigorous oversight reassures stakeholders that sensitive intellectual property is meticulously defended against unauthorized access while maintaining the collaborative transparency essential for modern multidisciplinary research.
Retrieval efficiency for legacy experiments through semantic tagging and full-text search capabilities
Unlocking decades of dormant laboratory wisdom requires sophisticated indexing mechanics that go far beyond basic matching to truly understand the contextual nuances of chemical research. Legacy paper archives often trap valuable experimental failures and unexpected side-reactions in dusty binders, making them virtually invisible to incoming generations of scientists who might repeat costly mistakes. By employing advanced semantic tagging architectures, electronic platforms automatically categorize reagents, catalyst concentrations, temperature profiles, and solvent systems into a deeply interconnected web of chemical knowledge.
When a researcher searches for a specific cross-coupling reaction, the system parses natural language descriptions alongside structural SMILES strings to instantly surface relevant historical insights from archives spanning twenty years or more. Full-text search engines index every handwritten note digitized via optical character recognition, every PDF supplement, and every raw spectrometer output file simultaneously. This capability drastically reduces redundant experimentation, allowing junior chemists to query past project successes within milliseconds and accelerate the optimization phase of active pharmaceutical ingredient development.
Navigating through vast historical datasets becomes an intuitive experience when search algorithms interpret molecular topologies rather than just strings of text. For instance, querying a specific heterocyclic core automatically retrieves related synthesis routes even if the original author used non-standard nomenclature or shorthand abbreviations. Visualizing search results through interactive network graphs reveals hidden correlations between disparate research projects conducted across different continents.
Researchers observe immediate efficiency gains because they spend less time hunting through physical archives and more time analyzing predictive reaction outcomes. Furthermore, metadata standards ensure that every legacy entry includes standardized units of measurement, yield percentages, and purity grades, creating a harmonized institutional memory that continuously feeds machine learning models designed to predict novel reaction pathways.
Standard operating procedures for transitioning a synthetic chemistry research group from paper journals to cloud-hosted documentation
Transitioning an established synthetic chemistry research group from traditional paper notebooks to a sophisticated cloud-hosted electronic laboratory notebook demands a carefully orchestrated strategy to secure buy-in and maintain operational continuity. Moving away from decades of muscle memory requires clear protocols, comprehensive training workshops, and empathetic leadership that addresses common anxieties surrounding data privacy and software complexity. Successful migration initiatives follow structured phased rollouts, ensuring that bench scientists experience immediate productivity benefits rather than administrative burdens.
- Phase one involves conducting a comprehensive audit of all active legacy projects to determine which historical records require digitization versus archival storage.
- Phase two establishes standardized metadata templates and controlled vocabulary lists for chemical reagents, equipment identifiers, and analytical techniques to ensure cross-team consistency.
- Phase three implements a pilot program with a select sub-group of enthusiastic early adopters who test the cloud platform under active bench conditions for a thirty-day cycle.
- Phase four delivers mandatory, hands-on training sessions focusing on secure cryptographic sign-offs, intellectual property protocols, and semantic search best practices for all remaining laboratory personnel.
- Phase five decommissions physical paper journals entirely, officially transitioning the entire department to a paperless, cloud-native operational standard supported by 24/7 IT helpdesk assistance.
To visualize the physical transformation of a modern synthetic chemistry laboratory during this migration, picture a sunlit, state-of-the-art facility where fume hoods hum quietly as researchers operate dual-monitor touchscreen terminals instead of flipping through heavy, ink-stained binders. Sleek glass reaction setups bubbling with vibrant organic compounds sit adjacent to ergonomic computer stations displaying real-time chromatographic traces linked directly to cloud entries.
Senior principal investigators review digital audit trails on portable tablet devices while walking between analytical suites, approving reaction schemes with secure cryptographic signatures from anywhere in the world. The absence of cluttered paper stacks creates an environment of focused precision, where every chemical interaction is instantly captured, securely timestamped, and woven into the collective intelligence of the global research enterprise.
Data integrity is the bedrock of scientific reproducibility; when cryptographic timestamps lock an experimental yield into the cloud, intellectual property transforms into an everlasting, unalterable asset for future innovation.
Molecular visualization applications empower researchers to inspect three-dimensional atomic structures and render publication-quality graphics for scientific journals.: What Software Does Chemists Use

Source: chemcomp.com
Stepping beyond raw numerical data and abstract columns of coordinates, modern computational chemistry breathes tangible life into invisible molecular realms. Through advanced graphical interfaces, scientists no longer merely imagine the spatial arrangements of atoms; they walk through them, manipulate them, and witness the delicate choreography of chemical architecture unfolding on high-resolution displays. These software suites serve as the ultimate bridge between rigorous mathematical calculation and visual comprehension, transforming complex quantum mechanical outputs into striking, intuitive masterpieces of structural science that communicate breakthrough discoveries to the global academic community.
To faithfully represent the unseen microscopic universe, these applications employ sophisticated rendering algorithms capable of simulating light propagation, shadow casting, and depth perception across complex atomic topologies. Ray tracing engines calculate the exact trajectories of light photons as they interact with transparent molecular surfaces, creating stunningly realistic reflections and refractions that convey true spatial depth. Ambient occlusion techniques subtly darken crevices and crowded atomic neighborhoods, providing an immediate visual cue regarding structural density without requiring explicit depth measurements.
Structural analysis rendering techniques
Advanced graphical shaders and lighting models are meticulously calibrated to expose the hidden forces and spatial constraints governing macromolecular behavior. By translating invisible physical fields into vivid chromatic maps, researchers can instantly diagnose chemical stability and reactivity.
- Hydrogen bonding networks are illuminated through pulsating dashed cylinders or glowing translucent tubes, dynamically updating their geometric parameters as bond lengths and angles fluctuate during molecular dynamics simulations.
- Electrostatic potential surfaces are mapped using continuous temperature color scales, where brilliant crimson regions denote electron-rich nucleophilic zones and deep sapphire hues highlight electron-deficient electrophilic areas ready for ligand attack.
- Steric clashes and unfavorable atomic overlaps are rendered as vivid, solid red spikes or jagged mesh protrusions erupting directly between van der Waals surfaces, instantly drawing the investigator’s eye to mechanical strain points.
Mechanisms of real-time collaborative viewing sessions for remote crystallography teams
Breakthrough discoveries rarely happen in isolation, and the modern structural biology laboratory spans continents, uniting crystallographers, pharmacologists, and virologists in synchronized investigation. Real-time collaborative viewing architectures solve the monumental challenge of sharing massive macromolecular datasets—frequently containing hundreds of thousands of individual atom coordinates—across distributed geographical networks without inducing debilitating latency. At the core of these collaborative frameworks lies a hybrid client-server synchronization protocol optimized specifically for high-frequency spatial data streaming.
Instead of continuously transmitting entire multi-megabyte coordinate files over standard internet connections, the host application broadcasts a lightweight stream of delta transformations, camera orientation vectors, and active selection states. Each remote participant runs a localized rendering engine that processes these incoming metadata streams, ensuring that the local hardware handles the heavy lifting of polygon rasterization and screen refresh rates at a fluid sixty frames per second.
Network overhead is further minimized through adaptive level-of-detail algorithms that dynamically simplify peripheral atomic structures while maintaining full, uncompromised resolution in the immediate focal point shared by the team. Secure WebSocket connections encrypt all transmitted session parameters in transit, safeguarding unpublished drug candidate structures and proprietary macromolecular conformations from interception. Furthermore, integrated voice-over-IP channels and synchronized pointer avatars allow multiple researchers to point, click, and annotate specific residues simultaneously, mirroring the natural collaborative workflow of standing shoulder-to-shoulder around a physical workstation.
When crystallographers solve a novel protein-ligand complex derived from synchrotron beamline data, this synchronized telepresence infrastructure enables immediate, cross-continental peer review and interactive model building, drastically accelerating the path from raw diffraction data to peer-reviewed publication.
Visualization is not merely a tool for illustration; it is the primary cognitive engine through which chemical intuition is forged, tested, and validated.
To assist research groups in selecting the optimal software environment for their specific computational pipelines, the following comparative matrix Artikels the core attributes of leading visualization platforms currently deployed in professional laboratories worldwide.
Modern laboratories rely on specialized computational tools to decode complex molecular structures, yet true breakthroughs emerge when raw data transforms into actionable clarity. By integrating mindfuel.ai product features dashboards analytics , researchers now visualize intricate reaction pathways instantly. This seamless digital evolution redefines what software does chemists use, bridging advanced mathematical modeling with intuitive daily discovery.
| Visualization Engine | Supported File Formats | OS Compatibility | Hardware Acceleration Requirements |
|---|---|---|---|
| PyMOL | PDB, CIF, SDF, MOL2, XYZ | Windows, macOS, Linux | Dedicated GPU with OpenGL 2.0+ support |
| ChimeraX | PDB, mmCIF, EM, MRC, Cheshire | Windows, macOS, Linux | Modern GPU supporting OpenGL 3.3 or Vulkan |
| VMD | PDB, DCD, TRJ, GRO, CHARMM | Windows, macOS, Linux | High-performance GPU with robust OpenGL shading |
| Avogadro | CML, XYZ, PDB, Gaussian Output | Windows, macOS, Linux | Standard graphics processor with basic OpenGL |
Chromatographic and spectroscopic data processing suites extract meaningful chemical concentrations and structural elucidation metrics from complex raw instrument signals.
Modern analytical chemistry thrives on precision, transforming chaotic streams of raw electronic signals into crystal-clear molecular insights. When high-performance liquid chromatography or mass spectrometry instruments complete a run, they output vast arrays of electrical voltages and ion counts that are completely unreadable in their raw form. Advanced data processing software acts as the ultimate digital lens, bringing obscured chemical realities into sharp focus and allowing scientists to quantify trace impurities or confirm novel synthetic pathways with absolute confidence.
Inside high-performance liquid chromatography software environments, baseline drift and overlapping peaks present formidable barriers to accurate quantification. Baseline correction algorithms utilize polynomial fitting or asymmetric least squares to mathematically flatten wandering signal foundations caused by temperature fluctuations or mobile phase gradients. Once a stable baseline is established, peak deconvolution steps in to resolve co-eluting compounds. By applying mathematical functions like Gaussian or Lorentzian curves to overlapping chromatographic envelopes, the software accurately separates adjacent peaks, determining individual retention times and peak areas with remarkable fidelity.
Automated spectral matching workflows used in nuclear magnetic resonance and mass spectrometry interpretation
The journey from a raw analytical run to a definitive molecular structure often relies heavily on automated spectral matching workflows, transforming days of manual analysis into mere seconds of computational wizardry. When a nuclear magnetic resonance spectrometer or a high-resolution mass spectrometer records an unknown compound, the resulting data file immediately enters a sophisticated processing pipeline designed for high-throughput identification.
This digital machinery systematically dissects the complex web of chemical shifts, coupling constants, and fragmentation patterns, comparing them against colossal reference libraries populated by millions of verified empirical entries.
At the heart of this automated marvel is the multi-stage filtering algorithm. Initially, the software performs a rapid pre-screening of the raw mass-to-charge ratios or proton-carbon correlation maps to eliminate structurally impossible candidates. It then deploys advanced pattern recognition algorithms, calculating similarity scores using Euclidean distance metrics, dot-product cosine calculations, or specialized probabilistic scoring models like reverse search algorithms. These computational protocols evaluate not just the presence or absence of specific peaks, but their relative intensities and isotopic fine structures.
For instance, in gas chromatography-mass spectrometry metabolomics, an automated workflow might ingest a raw blood plasma sample, deconvolute hundreds of overlapping chromatographic peaks, extract pure electron ionization spectra, and match them against the National Institute of Standards and Technology database within a minute.
Beyond simple library lookups, modern suites incorporate heuristic rules and machine learning architectures to predict spectra for hypothetical structures when exact matches fail. If an unknown molecule possesses a novel functional group modification, deep learning models trained on millions of quantum chemical calculations can forecast its expected carbon-13 chemical shifts or collision-induced dissociation pathways. The software ranks these probabilistic matches, offering the chemist a curated shortlist of structurally plausible isomers complete with confidence intervals.
This seamless fusion of empirical database searching and predictive artificial intelligence accelerates natural product discovery, environmental contaminant screening, and pharmaceutical impurity profiling, empowering researchers to decode the molecular architecture of complex mixtures with unprecedented speed and analytical depth.
Digital signal processing inevitably introduces subtle mathematical anomalies that can distort analytical outcomes if left unmanaged. Recognizing these artifacts is essential for maintaining the integrity of quantitative assays and structural elucidations.
- Gibbs ringing appears as artificial oscillatory ripples adjacent to sharp spectral boundaries, typically triggered by the truncation of finite Fourier series during digital filtering.
- Aliasing introduces false, lower-frequency signals into the data stream when analog waveforms are sampled below the critical Nyquist frequency threshold.
- Phase distortion skews symmetrical peak shapes into asymmetrical profiles, directly compromising accurate integration and reliable chemical shift determinations in nuclear magnetic resonance spectra.
- Quantization noise emerges as an unavoidable byproduct when continuous analog electrical voltages are converted into discrete digital integers by the analog-to-digital converter.
Mathematical precision in data processing transforms raw instrument noise into definitive molecular truth.
Chemical reaction drawing utilities serve as the universal graphical language for drafting complex synthetic pathways and generating standardized IUPAC nomenclature.
In the quiet hum of the modern chemical laboratory, ideas transform from abstract mental models into tangible blueprints through the stroke of specialized digital pens. Behind every groundbreaking pharmaceutical synthesis and peer-reviewed journal article lies an intricate dance of vectors, pixels, and rigorous chemical logic. Modern reaction drawing software acts as the primary translator of molecular architecture, bridging the gap between imaginative hypothesis and verifiable physical reality.
Researchers across the globe rely on these robust graphical environments not merely to sketch structures, but to enforce the unyielding laws of chemical behavior directly onto the digital canvas, ensuring that every synthesized ring, chain, and functional group communicates an unambiguous scientific truth.
Beneath the sleek interface of contemporary structure editors lies a sophisticated matrix of mathematical algorithms designed to prevent physical impossibilities on screen. When a chemist attempts to forge an impossible bond angle or force an eclipsed conformation that violates spatial realities, the software intervenes instantly. Advanced geometric constraint engines continuously calculate van der Waals radii, ring strain energies, and hybridization states in real time.
If a researcher tries to draw a planar carbon atom with five single bonds or distorts a tetrahedral geometry beyond acceptable thermodynamic thresholds, the application triggers immediate stereochemical validation rules. This proactive safeguarding ensures that chiral centers maintain their designated wedge-and-hash configurations, preventing costly misinterpretations during downstream pharmacological testing and automated database indexing.
Vector-based graphics engine integrations for crisp scaling in printed monographs and digital presentations
The transition from a raw computational workspace to a polished publication requires an uncompromising level of visual fidelity that raster images simply cannot deliver. By anchoring every atom symbol, bond stroke, and reaction arrow to mathematical vectors rather than fixed pixel grids, modern chemical drawing suites empower authors to scale their molecular artwork infinitely. Whether a complex total synthesis scheme is compressed into a tiny two-column journal thumbnail or projected across a massive auditorium screen for a keynote presentation, the visual integrity remains remarkably sharp.
Edges never blur, curves maintain their silky smoothness, and subscripts retain their crisp legibility even under extreme magnification. This mathematical foundation relies on scalable vector graphics principles where coordinates, line weights, and font Artikels are rendered dynamically by the viewing device. For publishing houses managing massive manuscripts filled with multi-step retrosynthetic trees, this vector integration eliminates the frustrating artifacts and pixelation common in older graphic formats.
Chemists rely on advanced molecular modeling suites to decode complex molecular structures, yet balancing laboratory budgets requires precision tools like price book software to streamline reagent costs. By mastering these diverse digital applications, modern researchers seamlessly bridge breakthrough scientific discovery with efficient financial management.
Furthermore, rich metadata accompanies these scalable graphics, embedding direct chemical connection tables right into the file structure. This means a graphic is never just a pretty picture; it remains an interactive, searchable data object capable of being queried by chemical information systems years after publication. Readers examining a complex natural product synthesis can click directly on an intermediate structure within a digital PDF, and the underlying vector framework immediately feeds the chemical graph into local modeling software for further analysis.
To achieve pristine visual standards across diverse media formats, drawing applications utilize specific structural scaling protocols:
- Standardized grid systems automatically snap molecular skeletons to predefined Cartesian coordinates to maintain uniform spatial distribution.
- Resolution-independent typography rendering engines scale atomic labels proportionally alongside bond length adjustments.
- Dynamic line-weight modulation ensures that bond thickness remains perceptually balanced regardless of overall diagram reduction or enlargement.
- Color-space management profiles calibrate RGB and CMYK output values seamlessly, bridging the gap between glowing monitor displays and high-resolution printing presses.
Procedures for standardizing bond lengths, atom labeling conventions, and reaction arrow alignments within collaborative drawing templates
Collaboration across multi-institutional research consortia demands absolute uniformity in graphical representation to prevent miscommunication among team members. When synthetic chemists, computational modelers, and analytical technicians share reaction schemes, inconsistent visual styles can lead to costly errors in laboratory execution. Establishing strict formatting protocols ensures that every multi-step mechanism reads with the clarity of a native scientific dialect. Laboratories typically deploy master template files pre-configured with institutional graphic design standards, compelling all contributors to adhere to unified aesthetic and spatial rules from the moment a new document is opened.
Implementing these universal templates involves several systematic adjustments to the editing environment:
- Configure global preference settings to enforce a fixed bond length, typically set to a standard dimension like 14.2 millimeters or an equivalent scale tailored for specific journal styles.
- Apply automatic atom-label alignment rules to ensure heteroatoms such as nitrogen, oxygen, and sulfur correctly orient their implicit hydrogen atoms based on spatial symmetry.
- Utilize automated alignment and distribution tools to snap reaction arrows, equilibrium harpoons, and retrosynthetic transform symbols onto a centralized horizontal axis.
- Embed explicit reaction conditions—such as catalytic reagents, temperatures, and solvent systems—into standardized text frames positioned at fixed vertical offsets above and below the reaction arrows.
- Lock template styles and custom toolbar presets into a shared cloud repository, forcing automated compliance checks before any reaction scheme can be exported for manuscript submission.
Precision in chemical drafting is not merely an aesthetic choice; it is the foundational language that prevents misinterpretation and drives reproducible innovation across the global scientific community.
Chemical inventory management platforms ensure workplace safety compliance by monitoring hazardous substance quantities and generating mandatory regulatory safety documentation.
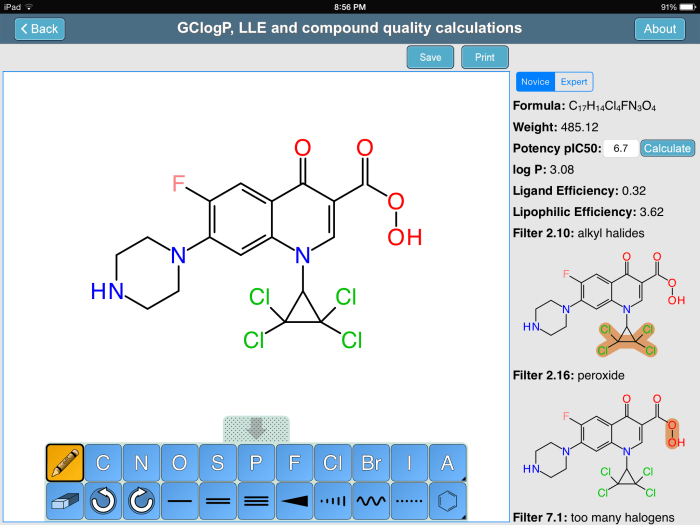
Source: cheminf20.org
Modern research facilities operate under strict safety guidelines where knowing the exact location and volume of every chemical container is vital for accident prevention. Digital stockroom solutions have revolutionized how laboratories handle hazardous materials, transforming chaotic physical shelves into streamlined, transparent digital ecosystems that prioritize human safety and ecological protection.
Behind the sleek interfaces of these safety platforms lies a sophisticated network of automated tracking tools designed to eliminate human error regarding volatile compounds. Chemical stockrooms house numerous substances that degrade over time into dangerously unstable states, requiring vigilant oversight that automated systems provide seamlessly without manual intervention.
Automated threshold alerting mechanisms for tracking expiration dates of peroxide-forming reagents and unstable compounds
Laboratories frequently store ethers, alkenes, and other peroxide-forming chemicals that silently build explosive crystalline shock-sensitivities as they age. Automated inventory suites continuously monitor elapsed time from the initial date of container opening, applying strict countdown algorithms tailored to specific chemical families. When a threshold approaches within thirty days, the platform dispatches high-priority notifications to safety officers and primary researchers, initiating mandatory testing protocols or safe disposal procedures.
Visualizing this monitoring architecture reveals a dynamic dashboard bathed in amber and crimson warning indicators, where virtual container icons pulse softly to demand immediate attention before physical degradation poses a genuine workplace hazard. These digital sentinels cross-reference environmental storage conditions with baseline chemical stability data, ensuring that volatile compounds never quietly slip past their safe usage windows into catastrophic instability.
Proactive shelf-life tracking transforms reactive emergency responses into predictable, routine maintenance schedules that protect laboratory personnel from unforeseen chemical degradation perils.
Integration processes linking chemical stockrooms with global Harmonized System hazard classification databases
Bridging local laboratory inventory software with international safety frameworks requires a robust, multi-layered digital infrastructure capable of translating localized stock data into universally understood hazard profiles. When a new chemical container enters a research facility and is scanned into the inventory platform, the system immediately initiates an API query against authoritative global repositories such as the United Nations Globally Harmonized System of Classification and Labelling of Chemicals.
Chemists decode molecular mysteries daily using advanced modeling software, transforming raw lab concepts into scalable industry reality through chemical manufacturing erp systems that streamline production pipelines, ultimately proving how cutting-edge digital tools empower scientists to accelerate global innovation safely.
This background synchronization extracts standardized pictograms, signal words, and hazard statements directly from official repositories, embedding them into the local database without manual data entry bottlenecks. The integration engine maps proprietary chemical CAS numbers and IUPAC nomenclature straight to international standards, ensuring that every digital record reflects the most current toxicological and ecological research available worldwide. Furthermore, as international safety committees update toxicity thresholds or reclassify specific compounds, the platform automatically propagates these updates across all active inventory logs, instantly flagging affected containers that may require updated secondary labeling.
This seamless data pipeline eliminates the dangerous discrepancy between local stockroom records and global safety mandates, providing regulatory inspectors with a cohesive, unassailable audit trail. Researchers benefit by accessing real-time safety data sheets directly from their mobile devices while standing inside the laboratory, bridging the gap between theoretical chemical knowledge and practical physical containment strategies. Ultimately, this global synchronization creates a unified defensive barrier, ensuring that regardless of where a compound is synthesized or stored, its inherent hazards are universally communicated, understood, and mitigated according to the highest international standards of occupational safety and environmental stewardship.
The following structured matrix Artikels the operational parameters required to maintain comprehensive safety compliance across modern research facilities:
| Regulatory Compliance Standards | Audit Frequency Schedules | Waste Disposal Tracking Methods | Emergency Response Data Integrations |
|---|---|---|---|
| OSHA Hazard Communication Standard | Quarterly internal physical verification | Manifest generation via EPA Uniform Hazardous Waste protocols | Real-time chemical location maps for local fire departments |
| EPA Resource Conservation and Recovery Act | Bi-annual comprehensive environmental audit | Barcode-scanned chain of custody tracking to licensed incinerators | Automated SDS transmission during hazardous materials spill events |
| NFPA 45 Fire Protection for Laboratories | Monthly inspection of storage cabinet integrity | Volume-based threshold monitoring for accumulation point limits | Direct API feeds to local hazardous materials response units |
Chemical safety and compliance software packages automate the generation of hazard communication labels and assess toxicological risks for new synthetic routes.
Modern laboratories operate on the razor-thin margin between groundbreaking innovation and catastrophic hazard, making safety intelligence an absolute priority for every scientific enterprise. Advanced digital platforms step into this high-stakes environment to shield researchers from unseen dangers, transforming raw molecular blueprints into clear, compliant action plans before a single reagent hits a beaker. By weaving regulatory oversight directly into the design phase, these tools redefine how chemistry protects its practitioners.
At the heart of these protective systems lie predictive toxicology modules that fundamentally change how scientists evaluate unknown hazards. When researchers sketch a novel compound, these algorithmic engines immediately cross-reference the atomic topology against massive global databases of known toxins. Using sophisticated machine learning classifiers and structural analogue matching, the software estimates critical safety benchmarks, such as the median lethal dose, known as LD50, with remarkable statistical confidence.
A vivid mental picture of this process reveals a digital workspace where a newly synthesized organic molecule glows on screen, instantly wrapped in a cascading waterfall of safety warnings, risk quotients, and toxicity probabilities derived from structural neighbors like benzene or aniline derivatives. This computational foresight allows teams to discard perilous synthetic routes on screen, sparing human hands from unnecessary exposure to volatile intermediates.
Predictive environmental fate modeling parameters for industrial chemical discharges
Understanding how a chemical behaves once it leaves the laboratory bench requires a deep dive into environmental fate modeling, a computational discipline that tracks a substance across air, water, and soil matrices. When industrial facilities prepare for large-scale production, compliance software calculates partition coefficients—such as the octanol-water partition coefficient, designated as K_ow—to determine whether a compound will bioaccumulate in aquatic food webs or remain dissolved in aqueous runoff.
Atmospheric degradation is simultaneously simulated by evaluating the molecule’s reaction rate constants with hydroxyl radicals, providing a clear picture of its tropospheric half-life and potential to contribute to ozone depletion or photochemical smog formation. Biodegradation kinetics are factored in through quantitative structure-activity relationships, which estimate the rate at which microbial communities in wastewater treatment plants can mineralize the pollutant into harmless byproducts like carbon dioxide and water.
Furthermore, soil sorption coefficients are meticulously derived to predict whether a chemical discharge will bind tightly to organic matter in sediment or leach rapidly through the soil profile into underlying groundwater aquifers. These complex calculations incorporate Henry’s law constants to forecast volatilization rates from liquid surfaces into the gas phase, ensuring that vapor emissions are accurately quantified under varying meteorological conditions.
Regulatory toxicologists rely heavily on these integrated mathematical simulations to establish safe discharge limits, design containment protocols, and verify that industrial processes will not trigger long-term ecological degradation across surrounding ecosystems. Through this rigorous computational lens, environmental stewardship becomes an exact science, anchoring industrial chemistry firmly to the carrying capacity of our planet.
Safety officers must implement stringent protocols when handling engineered particles whose nanoscale dimensions radically alter their biological interactions compared to bulk materials. The following measures Artikel the necessary protocols required for evaluating nanomaterial toxicity within enclosed laboratory containment facilities.
- Aerosol containment verification through high-efficiency particulate air filtration testing to prevent inhalation risks during nanopowder synthesis.
- Dynamic light scattering analysis of aerosolized working solutions to monitor real-time agglomeration states inside sealed gloveboxes.
- Cellular viability assays utilizing macrophage cultures exposed to standardized nanoparticle suspensions to gauge immediate cytotoxic responses.
- Surface reactivity profiling via electron paramagnetic resonance spectroscopy to quantify reactive oxygen species generation on particle interfaces.
Safety is not merely the absence of accidents, but the calculated elimination of hazard through digital foresight and uncompromising regulatory adherence.
Final Summary
Source: manufacturingchemist.com
Ultimately, the digital tools powering modern laboratories are far more than mere utilities; they are the true architects of tomorrow’s scientific breakthroughs. By seamlessly fusing rigorous data management with boundless computational creativity, these platforms elevate the entire research journey from a series of isolated experiments into a cohesive, visionary pursuit of knowledge. As technology continues to evolve, embracing these sophisticated environments ensures that the scientific community remains ever-ready to decode the universe’s most complex chemical mysteries with confidence, clarity, and enduring impact.
